DNA Methylation: Mechanism, Genomic Context and High-Throughput Analysis
Chemical Basis and Enzymatic Machinery of DNA Methylation
The DNMT family and 5-methylcytosine
In animals, DNA methylation is established and maintained by DNA methyltransferases (DNMTs):
-
DNMT1 – “maintenance” methyltransferase copying CpG methylation during DNA replication.
-
DNMT3A / DNMT3B – “de novo” methyltransferases that methylate previously unmethylated CpG sites.
-
DNMT3L – catalytically inactive regulatory factor in germ cells.
These enzymes transfer a methyl group from S-adenosyl-L-methionine (SAM) to cytosine C5 in the major groove. Mechanistic and structural details are summarized in reviews available via NCBI / PubMed on DNA methylation and epigenetic regulation (search “DNA methylation DNMT1 DNMT3”).
Comparative analyses across eukaryotes show that CG methylation is broadly conserved, and that Dnmt1-like maintenance methyltransferases retain similar catalytic cores in animals and plants. A classic cross-kingdom study is Feng et al., accessible via the UCSF library link to “Conservation and divergence of methylation patterning in plants and animals.” search.library.ucsf.edu

TET enzymes and cytosine oxidation
DNA methylation is reversible. In vertebrates, TET dioxygenases (TET1, TET2, TET3) oxidize 5mC to:
-
5-hydroxymethylcytosine (5hmC)
-
5-formylcytosine (5fC)
-
5-carboxylcytosine (5caC)
These oxidized bases are then removed by the base-excision repair pathway, leading back to unmethylated cytosine. Mechanistic descriptions of TET-mediated demethylation and base-resolution mapping of 5hmC can be found in open articles at NCBI PMC on oxidative bisulfite sequencing and 5hmC mapping. Academia
Passive demethylation during replication
Passive DNA demethylation occurs when DNMT1 is inhibited or absent during DNA replication. Newly synthesized strands remain unmethylated, and methylation density decreases with each cell cycle. This replication-coupled dilution is important in pre-implantation embryos and primordial germ cells, as summarized in reprogramming reviews available in NIH Roadmap Epigenomics perspectives at commonfund.nih.gov/epigenomics. commonfund.nih.gov+1
Sequence Context and Genomic Distribution
CG vs CHG vs CHH methylation
DNA methylation is often classified by sequence context:
-
CG methylation – predominant in vertebrates; symmetric (CpG on both strands).
-
CHG methylation – common in plants, symmetric, maintained by plant-specific CMT3 enzymes.
-
CHH methylation – asymmetric site; heavily used in plants for transposon silencing.
Reviews from Purdue University (“Dynamics and function of DNA methylation in plants,” PDF hosted at purdue.edu) and the Washington University in St. Louis biology department news (biology.wustl.edu) provide detailed descriptions of CG/CHG/CHH methylation and the associated methyltransferases in Arabidopsis and other plant species. Department of Biology+1
CpG islands, shores and shelves in vertebrate genomes
In mammalian genomes:
-
CpG islands are short (typically 300–3,000 bp), GC-rich regions enriched for CpG dinucleotides, often overlapping promoters and first exons.
-
CpG shores (0–2 kb from islands) and CpG shelves (2–4 kb from islands) are frequent sites for differential methylation in epigenome-wide association studies (EWAS).
Classical work on CpG islands, their features and relation to transcription is summarized in Deaton & Bird, accessible via Oxford Academic and often mirrored through institutional links (e.g. University of California or other .edu libraries). Academia
Promoter CpG islands of housekeeping genes are usually unmethylated, supporting open chromatin and transcription. In contrast, CpG islands associated with tissue-restricted or developmentally regulated genes may be methylated in non-expressing cell types.
Non-CpG (CpH) methylation in neurons and pluripotent cells
Non-CpG methylation (mCpH, H = A/C/T) is abundant in embryonic stem cells and post-mitotic neurons. It accumulates during neuronal maturation and is enriched in gene bodies and regulatory elements. Reviews and primary data on neuronal non-CpG methylation are available via NCBI PubMed and in open access at NCBI PMC (search “non-CpG methylation neurons 5hmC”).
These studies show that non-CpG methylation is recognized by MeCP2 and other methyl-binding proteins and contributes to fine-tuning of gene expression in the nervous system.
Organismal Diversity of DNA Methylation
Vertebrates and invertebrates
Comparative methylome studies show that:
-
Vertebrates typically have high global CpG methylation with unmethylated CpG islands.
-
Some invertebrates (e.g. honeybee, sea squirt) have methylation targeted mainly to gene bodies.
-
Other model organisms (e.g. Saccharomyces cerevisiae, Caenorhabditis elegans) have almost no detectable 5mC in genomic DNA. search.library.ucsf.edu
These observations are important for SEO phrases like “DNA methylation evolution”, “gene body methylation” and “epigenetic diversity in eukaryotes”.
DNA methylation in plants (CG, CHG, CHH)
Plant genomes show extensive methylation in CG, CHG and CHH contexts:
-
CG methylation in gene bodies and some promoters.
-
CHG / CHH methylation enriched in transposable elements (TEs) and repeats.
-
Distinct plant-specific methyltransferases such as CMT3 and DRM2 maintain non-CG methylation.
Comprehensive reviews on plant DNA methylation and its dynamics are available in PDFs hosted at Purdue University and other academic sites, such as:
-
“Dynamics and function of DNA methylation in plants” (Zhang et al.) purdue.edu
-
“Dynamics and malleability of plant DNA methylation during stress” (Lodhi et al., Thomas Jefferson University) jdc.jefferson.edu
These resources highlight how plant DNA methylation responds to abiotic and biotic stress, supports transposon silencing, and can be inherited across generations.
DNA methylation in transposable elements and repetitive DNA
Across eukaryotes, DNA methylation is a key mechanism for transposable element (TE) silencing. A detailed analysis of the epigenomic landscape of TEs across many tissues is provided by Pehrsson et al. in Nature Communications, available openly at NCBI PMC: “The epigenomic landscape of transposable elements across normal human tissues”. PMC
This work integrates DNA methylation, histone modifications and chromatin accessibility to show how TEs are differentially regulated in human cell types.
Functional Roles of DNA Methylation
Gene regulation and chromatin compaction
Key mechanisms linking DNA methylation to gene regulation:
-
Direct interference – 5mC at transcription factor binding sites can reduce DNA-protein binding.
-
Reader proteins – methyl-binding domain proteins (MBDs, MeCP2, etc.) bind methylated DNA and recruit histone deacetylases and chromatin remodelers, creating repressive chromatin.
-
Cross-talk with histone marks – DNA methylation interacts with histone modifications (e.g. H3K9me3) to stabilize heterochromatin.
These concepts are summarized in the NIH Roadmap Epigenomics Consortium overview and in reference epigenome analyses such as Kundaje et al., “Integrative analysis of 111 reference human epigenomes,” accessible at NCBI PMC. PMC+1
Development, imprinting and dosage regulation
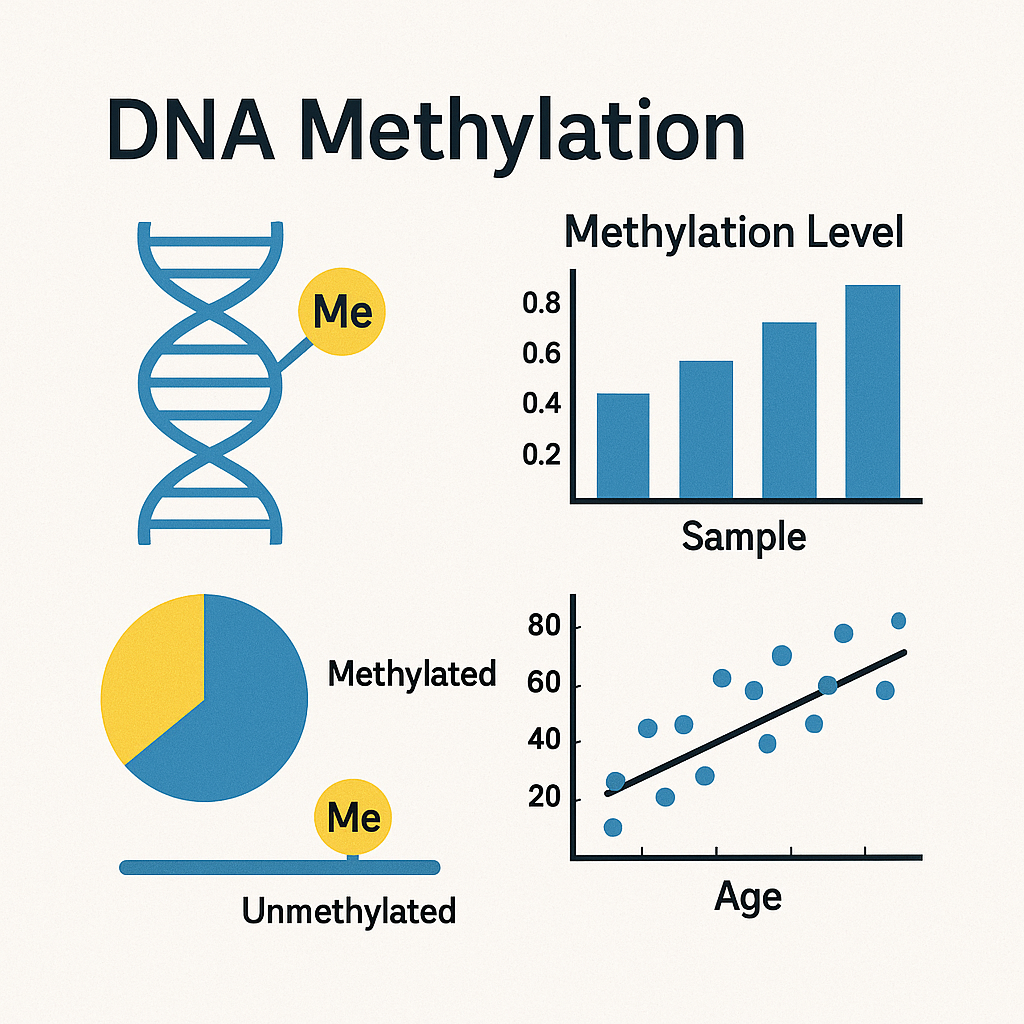
During early development, the genome undergoes global demethylation and remethylation, while specific loci retain methylation as imprints. These imprinted differentially methylated regions (DMRs) control parent-of-origin gene expression.
DNA methylation also contributes to:
-
X-chromosome inactivation in female mammals.
-
Stabilization of cell-type-specific transcription programs.
-
Prevention of aberrant activation of embryonic and germline genes in somatic tissues.
These functions are described in epigenetic reprogramming reviews linked through NIH and NCBI (search “DNA methylation dynamics epigenetic reprogramming”). PMC
Environmental responses and epigenetic variation
External factors—such as nutrient availability, temperature, stress and chemical exposures—can be associated with changes in DNA methylation patterns. In plants, stress-induced methylation changes are documented in reviews like Kumar et al. and Lodhi et al., focusing on abiotic stress and plant physiology, both accessible via academic and institutional repositories. librarysearch.chemeketa.edu+1
Population-level DNA methylation datasets raise questions about data interpretation, privacy and re-identification, discussed in policy-oriented work such as Dyke et al., “Epigenome data release: a participant-centered approach,” at NCBI PMC. PMC
Experimental Methods for DNA Methylation Analysis
Bisulfite conversion–based methods
Sodium bisulfite conversion is the gold standard for base-resolution DNA methylation analysis:
-
Unmethylated cytosine → uracil (C→T after PCR).
-
5mC remains as C.
Main techniques:
-
Whole-genome bisulfite sequencing (WGBS) – genome-wide CpG and non-CpG methylation at single-base resolution.
-
Reduced representation bisulfite sequencing (RRBS) – enriches CpG-rich regions for cost-efficient profiling.
-
Targeted bisulfite sequencing – captures selected loci or panels.
A practical WGBS tutorial is provided by UCLA QCBio in “Workshop 6 – WGBS,” PDF at qcb.ucla.edu. Academia
Workflows for plant and animal DNA methylation analysis with WGBS and RRBS, including adapter trimming and mapping, are reviewed in Omony et al. “DNA methylation analysis in plants: review of computational tools,” accessible via Oxford Academic / Briefings in Bioinformatics and mirrored in HTML on academic platforms. Academia
Array-based DNA methylation profiling (Illumina 450K and EPIC)
The Illumina HumanMethylation450 BeadChip (450K) and Infinium MethylationEPIC BeadChip are widely used for epigenome-wide association studies (EWAS). They measure methylation at hundreds of thousands of CpG sites.
Key methodological references (all open access at NCBI PMC, a .gov resource):
-
Okamura et al. – probe content and annotation of 450K. Academia
-
Lehne et al. – “A coherent approach for analysis of the Illumina 450K array.” Academia
-
Pidsley et al. – critical evaluation of the EPIC array. Academia
-
Kundaje et al. – integrative analysis of 111 reference epigenomes, including 450K data. PMC
These articles detail normalization, probe filtering, batch correction and differential methylation analysis, forming a standard pipeline for EWAS-style DNA methylation profiling.
Enrichment-based methods (MeDIP-seq, MBDCap-seq, Methyl-Capture)
Before WGBS became standard, many studies used enrichment of methylated DNA:
-
MeDIP-seq – immunoprecipitation using antibodies against 5mC.
-
MBDCap-seq – capture of methylated DNA using methyl-CpG-binding domains.
-
Methyl-Capture – hybridization-based enrichment of CpG-dense regions.
These methods provide regional methylation profiles at lower cost and can still be useful for screening large sample cohorts, as described in methodological reviews on NCBI PMC and in epigenomics resource guides. PMC
Direct detection with long-read sequencing
New long-read platforms enable direct detection of cytosine modifications:
-
PacBio HiFi “5-base sequencing” infers 5mC from polymerase kinetics; an application brief is hosted at UMass Chan Medical School: “Measuring DNA methylation with 5-base HiFi sequencing”. repositori.upf.edu
-
Oxford Nanopore devices detect methylation from changes in ionic current, enabling real-time DNA methylation sequencing of long molecules; overviews are provided in reviews on profiling genome-wide DNA methylation at NCBI PubMed / PMC. Academia
These methods support advanced SEO phrases like “long-read DNA methylation sequencing”, “5-base HiFi epigenome profiling”, and “nanopore DNA methylation assay”.
Single-Cell and Multi-Omic DNA Methylation
Single-cell DNA methylome sequencing
Single-cell bisulfite sequencing (scBS-seq) and related methods combine ultra-low-input bisulfite conversion with indexing strategies to profile DNA methylation at the single-cell level. These approaches capture cell-to-cell heterogeneity in CpG methylation and can be integrated with single-cell RNA-seq or single-cell ATAC-seq.
Many protocols and benchmarking studies are accessible via NCBI PMC and university sequencing cores (e.g. Northwestern University NUSeq – DNA methyl-seq services). Academia
Integration with chromatin accessibility and histone marks
ATAC-seq and DNA methylation can be combined to model chromatin accessibility and methylation state at regulatory elements. For example, Zhong et al. describe how DNA methylation-linked chromatin accessibility shapes transcription in plants, accessible as a PDF from UCLA at research.mcdb.ucla.edu. research.mcdb.ucla.edu
At the consortium level, NIH Roadmap Epigenomics and ENCODE provide integrated datasets of DNA methylation, histone modifications and DNase/ATAC-seq. Users can browse these data at:
-
The NIH Roadmap Epigenomics portal: commonfund.nih.gov/epigenomics commonfund.nih.gov
-
The ENCODE portal: encodeproject.org (described in Jou et al. at NCBI PubMed). PubMed
Public Epigenome Resources and Data Portals
NIH Roadmap Epigenomics Mapping Consortium
The NIH Roadmap Epigenomics Mapping Consortium generated large reference datasets of DNA methylation, histone marks and open chromatin across many human cell types and tissues. Its goals and design are described in:
-
The original consortium paper at NCBI PMC: “The NIH Roadmap Epigenomics Mapping Consortium”. PMC
-
The dedicated resource article “The NIH Roadmap Epigenomics Program data resource” at NCBI PMC. PMC
These datasets are accessible through:
-
Roadmap Epigenomics site (via commonfund.nih.gov/epigenomics). commonfund.nih.gov
-
The ENCODE portal. PubMed
International Human Epigenome Consortium (IHEC)
The International Human Epigenome Consortium (IHEC) coordinates international epigenome projects and exposes unified datasets through the IHEC Data Portal. The portal is described in “The International Human Epigenome Consortium Data Portal” at NCBI PubMed: pubmed.ncbi.nlm.nih.gov/27863956. PubMed
IHEC integrates data from ENCODE, NIH Roadmap, Blueprint, DEEP, and other consortia, enabling cross-study comparison of DNA methylation and related marks.
Epigenome browsers and visualization tools
A guide to epigenome browsers and data resources is provided by Karnik et al. in “A guide to data resources and epigenome browsers for human epigenome project data,” available at NCBI PMC: pmc.ncbi.nlm.nih.gov/articles/PMC3750740. PMC
This guide covers:
-
The UCSC Genome Browser (University of California, Santa Cruz – genome.ucsc.edu)
-
The WashU Epigenome Browser (Washington University in St. Louis – epigenomegateway.wustl.edu)
-
Specialized viewers for Roadmap and ENCODE datasets
These platforms host high-density tracks of DNA methylation, making them central for DNA methylation analysis, EWAS visualization, and regulatory annotation.
Computational Pipelines for DNA Methylation Data
Pre-processing of bisulfite sequencing
Typical computational steps for WGBS / RRBS:
-
Quality control and adapter trimming (e.g. Trimmomatic).
-
Bisulfite-aware alignment to the reference genome (e.g. Bismark, BS-Seeker, BS-Seeker3, WALT, GEMBS).
-
Methylation calling at each cytosine.
-
Aggregation of methylation proportions per CpG or region.
The review “DNA methylation analysis in plants: review of computational tools” lists many tools and their use cases, including BS-Seeker2/3, WALT, BiQ Analyzer, BSeQC, and DMR callers such as Metilene, DMRcaller and others. Academia
EWAS and differential methylation
In epigenome-wide association studies:
-
Beta values (methylated intensity / total) or M-values (logit transform) are used.
-
Linear models and mixed models capture associations with traits or exposures.
Statistical frameworks for differentially methylated loci (DML) and differentially methylated regions (DMR) are reviewed in detail in Omony et al. and in genome-wide epigenomics methods papers linked through NCBI PMC. Academia+1
Integration with genomic annotations
DNA methylation data are typically intersected with:
-
Promoters, enhancers and CpG islands (from UCSC or Ensembl).
-
ChromHMM/Segway chromatin states from ENCODE and Roadmap.
-
Transposable element annotations (e.g. from RepeatMasker tracks).
Epigenome browsers described by Karnik et al. provide built-in tools to overlay DNA methylation with these tracks. PMC
Experimental Design, Controls and Quality Metrics
Technical controls
Robust DNA methylation assays usually include:
-
Unmethylated control DNA (e.g. lambda phage DNA) to monitor bisulfite conversion efficiency.
-
Fully methylated control DNA to check assay sensitivity.
-
Sample replicates to estimate technical variance.
Guidelines on coverage recommendations and QC for WGBS are summarized in methodological reviews referenced in Omony et al. and related articles at NCBI PMC. Academia
Quality control metrics
Common QC metrics:
-
Conversion rate (fraction of non-CpG cytosines converted to T).
-
Global CpG methylation levels compared with expectations for given cell types.
-
Coverage distribution and CpG read depth.
-
Replicate concordance (e.g. Pearson correlation of methylation profiles).
Visualization and QC tools, such as BiQ Analyzer and BSeQC, are discussed in Omony et al. and related software articles (Bioinformatics journal). Academia
You can reuse this block directly in your Affigen / Gentaur / health.gentaur.com pages:
DNA Methylation – Epigenetic Cytosine Modification and Genome-Wide Profiling
DNA methylation is a key epigenetic modification in which methyl groups are added to cytosine residues, mainly at CpG sites, generating 5-methylcytosine. This covalent change does not alter the DNA sequence but has major effects on gene regulation, chromatin structure, genomic imprinting, X-chromosome inactivation, transposon silencing and long-term epigenetic memory. Modern DNA methylation analysis combines bisulfite conversion, whole-genome bisulfite sequencing (WGBS), reduced representation bisulfite sequencing (RRBS), targeted bisulfite assays, Illumina 450K and MethylationEPIC arrays, and long-read DNA methylation sequencing with advanced bioinformatics pipelines for differential methylation and epigenome-wide association studies (EWAS). Public epigenome resources such as the NIH Roadmap Epigenomics Program, ENCODE, and the International Human Epigenome Consortium (IHEC) provide thousands of reference DNA methylation maps across human tissues and cell types, enabling integrative analysis of CpG methylation, non-CpG methylation, chromatin accessibility and histone modifications.